Top 10 Phenylacetonitrile Intermediates Essential for Organic Synthesis
2026-10-06
Phenylacetonitrile intermediates quietly power a surprising share of modern organic chemistry—yet most chemists only ever reach for the same two or three familiar building blocks. In this post, we break down the top 10 phenylacetonitrile derivatives that deserve a spot in every synthetic toolbox. From streamlining heterocycle construction to enabling late-stage functionalization, these workhorse compounds solve real problems at the bench. Whether you're optimizing a route or chasing a novel scaffold, the right nitrile can cut steps, boost yields, and open new disconnections. Drawing on hands-on experience from DSL Chemicals, we've ranked each intermediate by practical value, not just literature popularity. Read on to see which underused derivatives could transform your next synthesis.
Phenylacetonitrile as a Launchpad for Heterocyclic Scaffolds
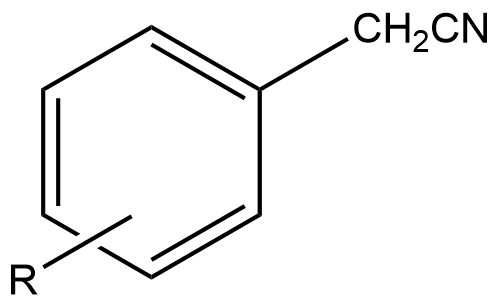
The benzylic position of phenylacetonitrile offers a reactive methylene group that can be deprotonated with mild bases, generating a nucleophilic carbanion. This intermediate readily participates in alkylations, acylations, and Michael additions, installing diverse side chains before the nitrile is engaged in ring-forming steps. Because the phenyl ring remains intact through many transformations, it provides a useful handle for further functionalization or can be removed later if needed.
The nitrile itself is far from a spectator. Under acidic, basic, or metal-catalyzed conditions, it can be converted into thioamides, amidines, or carboxylic acid derivatives, which then serve as direct precursors to heterocycles. For instance, condensation with hydrazines yields aminopyrazoles, while reaction with hydroxylamine gives isoxazoles. Cyclization with ortho-functionalized aryl partners opens routes to quinolines, isoquinolines, and indoles, making phenylacetonitrile a compact entry point to fused ring systems.
What makes this scaffold particularly attractive is the ability to orchestrate sequential C–C and C–N bond formation without isolating sensitive intermediates. A single pot can take phenylacetonitrile through deprotonation, electrophilic trapping, and nitrile-directed cyclization, often with good regiocontrol. This efficiency, combined with the low cost of the starting material, has kept it relevant in medicinal chemistry campaigns and natural product-inspired synthesis.
Reductive Transformations to Phenethylamine-Type Intermediates
The conversion of substituted nitrostyrenes into phenethylamine-type intermediates often hinges on the choice of reducing agent and the control of competing pathways. Lithium aluminum hydride remains a classic option for fully reducing the nitroalkene to the saturated amine, but its harsh profile can complicate substrates bearing sensitive esters or halides. Milder borohydride systems, particularly when paired with transition metal salts, allow stepwise reduction through the hydroxylamine or oxime stage, which is useful when the goal is to isolate intermediates rather than rush directly to the primary amine.
Catalytic hydrogenation offers another route, though the selectivity profile shifts with catalyst and pressure. Palladium on carbon tends to deliver the saturated phenethylamine cleanly when the alkene is conjugated to the nitro group, but over-reduction of aromatic rings or dehalogenation can occur if conditions are not tightly managed. Platinum or Raney nickel catalysts sometimes preserve the aromatic substituents better, but they may require acidic additives to prevent dimerization of the intermediate hydroxylamine species.
A less obvious but increasingly practical approach uses dissolving metal reductions, such as zinc or iron in acetic acid, which generate the amine directly from the nitroalkene while tolerating certain carbonyl groups. These methods often produce fewer byproducts from alkene migration because the reduction proceeds through a radical or single-electron pathway rather than a clean hydride addition. Selecting the right combination of metal, acid strength, and temperature can shift the outcome toward either the fully saturated phenethylamine or the partially reduced oxime, giving chemists room to tune the intermediate for downstream functionalization.
Hydrolysis and Amidation Routes to Phenylacetic Acid Derivatives
Transforming phenylacetic acid into its derivatives often starts with controlled hydrolysis of precursor esters or nitriles. Alkaline hydrolysis of benzyl cyanide, for instance, gives the sodium salt of phenylacetic acid, which can be acidified to the free acid without isolating intermediates. This route benefits from mild temperatures and avoids over-oxidation, making it a practical entry point when the aromatic ring carries sensitive substituents. By adjusting the base strength and reaction time, one can limit decarboxylation and preserve the integrity of the side chain.
Amidation, on the other hand, converts phenylacetic acid into amides that serve as key intermediates in pharmaceuticals and agrochemicals. A common method involves activating the carboxylic acid with carbodiimides or converting it to the acid chloride, then reacting with primary or secondary amines. For less reactive substrates, mixed anhydrides or coupling agents like HATU offer better yields under ambient conditions. Interestingly, direct thermal amidation with amine salts can avoid auxiliary reagents altogether, though it requires higher temperatures and careful removal of water. This route is often preferred for large-scale work because of lower cost and simpler purification.
Combining hydrolysis and amidation in a single sequence allows chemists to build diverse libraries from a common phenylacetic acid scaffold. For example, an ester is first hydrolyzed to the acid, then immediately coupled to a chiral amine under mild conditions to preserve stereochemistry. Choosing the right solvent and pH during the hydrolysis step can suppress racemization when dealing with alpha-substituted analogues. Tougher cases, where the phenyl ring bears electron-withdrawing groups, may need enzymatic hydrolysis to avoid harsh reagents. Ultimately, the choice between chemical and enzymatic methods hinges on selectivity, scale, and the tolerance of downstream functional groups.
Alpha-Carbon Chemistry: Alkylation and Arylation Patterns
Alkylation at the alpha carbon usually hinges on how well the enolate forms and how stable it remains before the electrophile arrives. With a ketone or aldehyde, a slight excess of strong base like LDA in a polar aprotic solvent can generate the kinetic enolate, which then reacts with primary alkyl halides or tosylates to give predictable substitution. The pattern changes when the electrophile is bulky or when the substrate can form two different enolates; here, temperature and base choice steer the product toward either the less substituted or more substituted alpha position.
Arylation introduces a different electronic demand because aryl halides are reluctant to undergo direct SN2 displacement at an sp2 carbon. Instead, palladium or copper catalysis often links the enolate or its equivalent to an aryl group. For example, the alpha-arylation of ketones with aryl bromides or chlorides can be achieved with a palladium catalyst and a bulky phosphine ligand, allowing even hindered substrates to couple cleanly. The reaction tolerates electron-rich and electron-poor aryl halides, though ortho-substituted aryl halides may require higher temperatures or modified ligands to avoid sluggish oxidative addition.
Malonic ester and acetoacetic ester syntheses remain the classic teaching examples because the alpha carbon between two carbonyls is acidic enough to be alkylated under mild basic conditions. After monoalkylation, a second alkylation can be performed before hydrolysis and decarboxylation, which makes these routes efficient for building quaternary centers or substituted acetic acids. In arylation variants, the same dicarbonyl frameworks can be coupled with aryl iodides under copper catalysis, then decarboxylated to yield alpha-aryl acetic acid derivatives without resorting to harsh direct arylation conditions.
Halogenated Phenylacetonitriles in Modern Cross-Coupling
Halogenated phenylacetonitriles occupy a curious niche in cross-coupling chemistry. The electron-withdrawing nitrile group, particularly when positioned ortho or para to the halogen, exerts a strong influence on the aryl ring's electron density. This alters the rates of oxidative addition and transmetallation in ways that are not always intuitive. For instance, 2-bromophenylacetonitrile often undergoes Suzuki-Miyaura coupling more sluggishly than its unsubstituted counterpart, yet the same substrate can be coaxed into efficient reactions with careful selection of bulky, electron-rich phosphine ligands. The nitrile also serves as a versatile handle for post-coupling transformations, which makes these building blocks attractive despite their sometimes frustrating reactivity.
In practice, the synthetic value of halogenated phenylacetonitriles becomes apparent in heterocycle synthesis. A common strategy involves a palladium-catalyzed Heck reaction between an ortho-halogenated phenylacetonitrile and an activated alkene, followed by intramolecular cyclization to afford indanone or isoquinoline derivatives. The nitrile group can act as a directing group in some C-H activation scenarios, though its weak coordination often requires finely tuned reaction conditions. Negishi couplings with organozinc reagents derived from these substrates have also been reported, especially for the construction of biaryl acetonitriles that would be difficult to access via classical routes. The key is often not the choice of metal but the ligand architecture and the base, which must balance the nitrile's tolerance against the need for efficient turnover.
Recent advances have pushed these substrates into less conventional territory. Dual photoredox-nickel catalysis has enabled the coupling of halogenated phenylacetonitriles with alkyl radicals under mild, visible-light-driven conditions, avoiding the need for strong external bases that could hydrolyze the nitrile. Electrochemical methods have similarly allowed for reductive cross-coupling without stoichiometric metal reductants. Continuous flow setups have also improved the scalability of these reactions by mitigating exotherms and enhancing mixing, which is particularly helpful for challenging substrates like ortho-substituted phenylacetonitriles. As mechanistic understanding deepens, the once-troublesome reactivity of these compounds is increasingly seen as a tunable feature rather than a limitation.
Nitrile-Directed Metalation for Ring Functionalization
The nitrile group, often dismissed as a mere spectator in aromatic chemistry, turns into a powerful steering ligand when paired with strong bases. Its lone pair on nitrogen coordinates transiently to alkali metal amides, pulling the metal to the ortho position even when steric or electronic factors would normally send it elsewhere. This subtle tug reorders the usual reactivity map, enabling selective deprotonation at a ring carbon that conventional directing groups struggle to reach.
Once the metal sits ortho to the cyano substituent, the resulting arylmetal intermediate becomes a versatile handle. Electrophiles ranging from iodine to aldehydes can be introduced with predictable regiochemistry, while the nitrile itself remains untouched. For substrates bearing multiple aromatic rings, this method sidesteps the need for protecting groups or prefunctionalized halides, offering a direct route to densely substituted benzonitriles that would otherwise require lengthy detours.
In practice, the reaction’s success hinges on matching the base and solvent to the substrate’s acidity. Lithium dialkylamides in ethereal solvents strike a balance between deprotonation strength and coordination lifetime, whereas bulkier amide bases can override the nitrile’s directing effect when competing coordination sites exist. Temperature control further fine-tunes the selectivity, making this strategy a go-to for medicinal chemists who need late-stage diversification of cyanoarene scaffolds without disturbing sensitive functionality elsewhere in the molecule.
FAQ
The nitrile acts as a masked carboxylic acid. It tolerates many conditions that would degrade an ester or acid, yet it can be hydrolyzed to phenylacetic acid or converted to an amide or amine depending on the reagent. That switchable behavior lets chemists carry the phenylacetonitrile unit through multiple steps before revealing the final functional group.
Phenylacetic acid, phenylacetamide, 2-phenylethylamine, and phenylacetone are the workhorses. Phenylacetic acid is used to make penicillin derivatives, phenylacetamide appears in some anticonvulsant routes, 2-phenylethylamine leads to sympathomimetic amines, and phenylacetone is a known precursor for amphetamine-class compounds.
Controlled hydrolysis with hydrogen peroxide and a mild base, or brief treatment with concentrated sulfuric acid at room temperature, gives the amide selectively. The key is to stop the reaction once the nitrile has added water once, because prolonged heating pushes it to phenylacetic acid.
Catalytic hydrogenation with Raney nickel in ethanol containing ammonia is the standard choice. Adding ammonia suppresses coupling to secondary and tertiary amines, so the primary amine is isolated in high yield after filtering off the catalyst.
First hydrolyze phenylacetonitrile to phenylacetic acid, then heat the acid with acetic anhydride and a sodium acetate catalyst. The reaction forms a mixed anhydride that decarboxylates, and fractional distillation gives phenylacetone clean enough for further use.
Yes. Treating it with a strong base like sodium amide or LDA generates a stabilized anion that can be quenched with alkyl halides. Mono-alkylated phenylacetonitriles, such as 2-phenylbutyronitrile, are valuable precursors to branched phenylacetic acids and amines used in specialty pharmaceuticals.
2-Phenylpropionitrile, made by methylating phenylacetonitrile, is a common precursor to 2-phenylpropionic acid. Enzymatic or chemical resolution of this nitrile gives enantiomerically enriched material, which fits into a family of arylpropionic acid scaffolds used in medicinal chemistry.
The nitrile route avoids the lachrymatory and genotoxic issues associated with benzyl chloride. Phenylacetonitrile is easier to handle, has better stability in storage, and its reactions usually generate fewer problematic byproducts, which simplifies scale-up.
Conclusion
Phenylacetonitrile has long served as a compact, bifunctional starting material whose benzylic C–H and nitrile carbon invite independent manipulation. Among the most valuable intermediates in this family are those used to assemble heterocyclic cores. Condensation with hydrazines, amidoximes, or guanidine equivalents converts the nitrile into pyrazoles, oxadiazoles, and aminopyrimidines, often in two or three steps. Reductive processing of the same intermediate stream yields phenethylamine-type building blocks; by stopping at the aldimine or amidine stage, chemists preserve a partially activated nitrogen that can be carried into reductive amination or cyclization. Hydrolysis routes are equally practical: acid- or base-promoted hydration gives phenylacetamide, while full hydrolysis leads to phenylacetic acid. These three product classes—heterocycles, amines, and carboxylic acid derivatives—account for many of the top-ten intermediates because they preserve the aromatic ring for later modification.
The remaining high-value intermediates arise from selective modification of either the α-carbon or the aryl ring. Deprotonation at the benzylic position allows alkylation with primary and secondary halides, arylation with aryl halides under palladium or copper catalysis, and sequential double substitution to create quaternary centers; the resulting branched phenylacetonitriles are direct precursors to profen-class analgesics and tertiary carbinamine pharmacophores. Halogenated phenylacetonitriles, particularly 2-, 3-, and 4-bromo or chloro derivatives, have become reliable partners in Suzuki–Miyaura, Buchwald–Hartwig, Sonogashira, and Ullmann couplings. Because the nitrile remains intact under these conditions, the coupled biaryl or alkyne products can still undergo nitrile hydrolysis or reduction. In parallel, directed ortho-metalation of phenylacetonitrile—usually with lithium amide bases or TMP-zincate reagents—places boron, silicon, halogen, or thioether groups at the ring positions adjacent to the acetonitrile side chain. This combination of ring and side-chain handles makes the top phenylacetonitrile intermediates indispensable for convergent synthesis of drug-like scaffolds.
Contact Us
Contact Person: Wei Zhang
Email: [email protected]
Tel/WhatsApp: 862163529955
Website: https://www.dslchem.com